
MinerAlert
MinerAlert
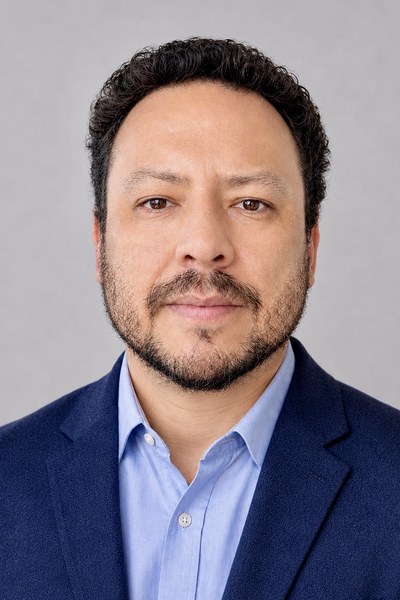
Jesús Naín Pedroza-Montero, Ph.D.
Interdisciplinary Research Building (IDRB)
Department: Physics
Dr. Jesús Naín Pedroza-Montero is an Assistant Research Professor at The University of Texas at El Paso. His research is in computational quantum chemistry and electronic structure methods, with an emphasis on the numerical algorithms and software that make density functional theory (DFT) calculations feasible for large molecular and nanoscale systems. He completed his doctoral studies in 2020 in the Nanosciences and Nanotechnology program at Cinvestav (Centro de Investigación y de Estudios Avanzados del Instituto Politécnico Nacional) in Mexico City, where he worked in the deMon2k development group led by Andreas M. Köster and Patrizia Calaminici.
Dr. Pedroza-Montero is a listed developer of deMon2k, a DFT program built on the auxiliary density functional theory (ADFT) framework. His publications include “Variational Density Fitting with a Krylov Subspace Method” (Journal of Chemical Theory and Computation, 2020), which introduced an iterative linear algebra approach to density fitting, and “Variational Fitting of the Fock Exchange Potential with Modified Cholesky Decomposition” (The Journal of Chemical Physics, 2020). He has also collaborated with the CNRS and Université Paris-Saclay group on real-time time-dependent ADFT simulations. His research interests include quantum chemistry, DFT, electronic structure methods, data science, and software development.
More recently, Dr. Pedroza-Montero has been collaborating with researchers at Pacific Northwest National Laboratory (PNNL) on the development and application of advanced computational approaches for large-scale molecular simulations. This work includes atom-specific vibrational analyses aimed at identifying labile bonds and structure-dependent reactivity in linear and branched perfluorooctanoic acid (PFOA) molecules, as well as the implementation of electronic-structure simulation workflows in cloud and high-performance computing environments. A growing component of these efforts focuses on the integration of machine-learning interatomic potentials with quantum-chemical methodologies to accelerate geometry optimizations, conformational exploration, and molecular property calculations for systems that are beyond the practical scale of conventional first-principles simulations. These machine-learning methodologies are also being extended to applications in energy-storage and battery materials, as well as to the phenomic analysis of complex biological systems, with the goal of enabling efficient, large-scale exploration of structure-property relationships across chemically and biologically diverse systems.
Keywords of Expertise: Computational Quantum Chemistry, Density Functional Theory, Electronic Structure Methods, Scientific Software Development, Numerical Linear Algebra, Data Science, Machine Learning Interatomic Potentials.